Compute overall (multi-sample) dissimilarity metrics from the Hill-number
beta diversity following Chiu et al. (2014). These are the complements of the
similarities returned by hillsim(). With hierarchy, each transition's
beta is converted using its data-dependent effective ceiling rather than
the number of child units, keeping the metrics in [0, 1].
Arguments
- data
A count table (taxa x samples) or a supported object; a single sample is not meaningful for partitioning.
- q
Numeric vector of diversity orders (>= 0). Defaults to
c(0, 1, 2)(richness, Shannon, Simpson).- metric
Dissimilarity metric(s) to return, any of
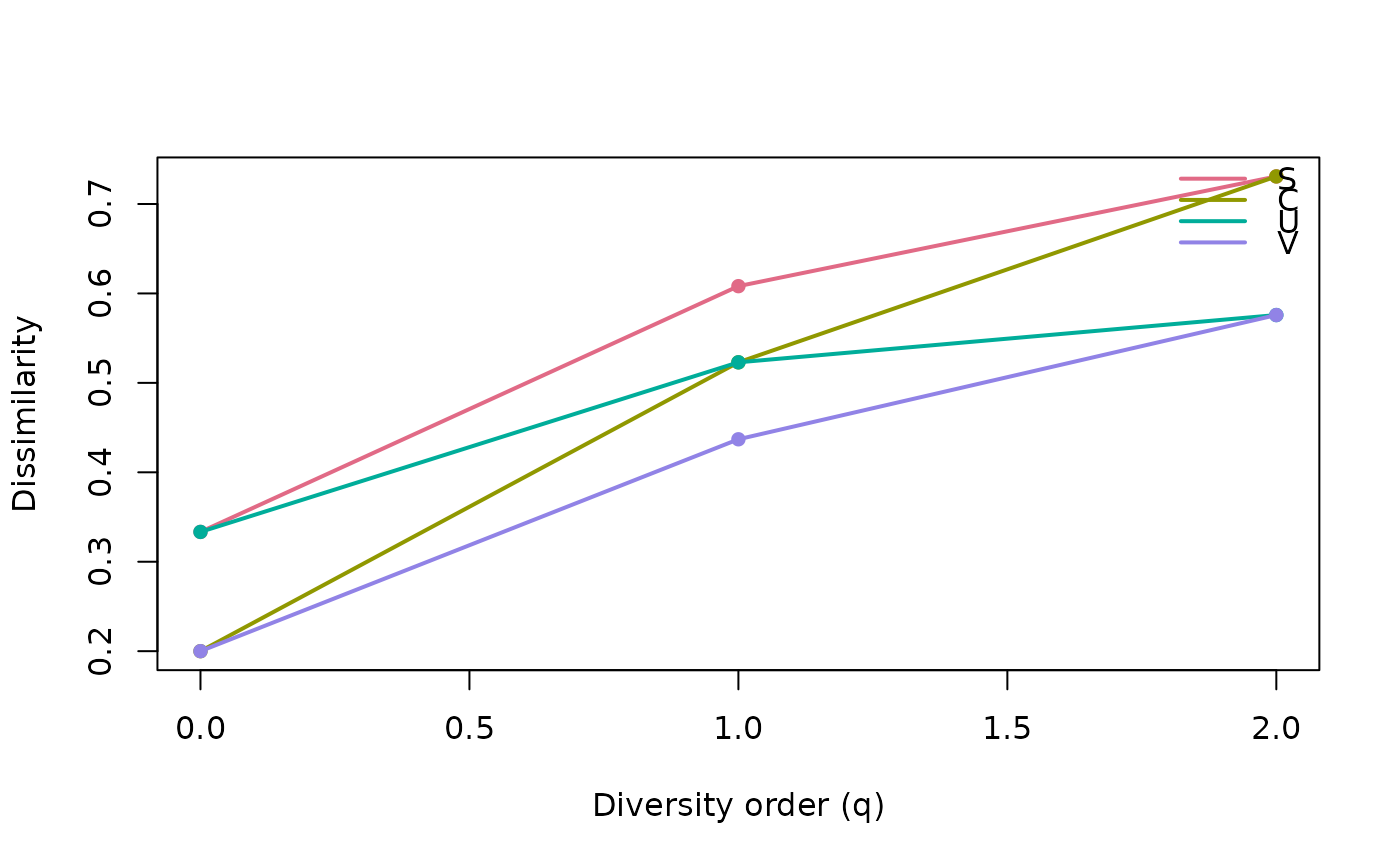
"S","C","U","V". Defaults to all four."S"(Jaccard-type) and"V"(Sorensen-type) are turnover measures, relative to the pooled assemblage and to a single assemblage respectively;"U"(Jaccard-type) and"C"(Sorensen-type; one minus the Morisita-Horn similarity atq = 2) are the corresponding overlap complements (proportions of non-shared units). Atq = 0,"U"equals"S"and"C"equals"V"; atq = 1,"U"equals"C".- tree
A phylogenetic tree of class
phylowhose tip labels match the taxa indata.- dist
A functional distance matrix (or
dist) over the taxa.- tau
Optional functional distance threshold. Defaults to
max(dist).- type
Diversity type:
"auto"(default) infers it from the inputs (counts only -> neutral,+tree-> phylogenetic,+dist-> functional); an explicit"neutral","phylogenetic"or"functional"asserts the type and is validated against the inputs (e.g."phylogenetic"requires atree;"neutral"ignores any tree/dist carried by the object).- out
Output shape:
"tibble"(default) returns a long-formatdata.framewith columnsq,metric,value;"matrix"returns the legacy matrix (orders in rows, metrics in columns, dropped to a vector for a single metric). Withhierarchy, the table adds ascalecolumn for each transition above the sample scale. Ataugrid adds ataucolumn. Matrix output has one row perqand transition, with a named list of matrices for ataugrid.- hierarchy
Optional one-sided nesting formula, coarsest to finest, e.g.
~ region / site. Every transition abovesample, includingtotal, gets its own dissimilarity metrics. Each run uses one shared treedepthor functionaltau; a functionaltaugrid runs the full hierarchy once per threshold.- metadata
Optional per-sample
data.framesupplying the variables named inhierarchy; rows are matched to the count-table columns by name when possible, otherwise by position.- weights
Sample weights for hierarchical partitioning:
"equal"(default),"abundance"(proportional to each sample's total count), or a positive numeric vector with one value per sample. A named vector is matched to count-table columns; an unnamed vector uses column order. Weights are normalised to sum to one. Requireshierarchy.- depth
Optional positive phylogenetic reference depth for hierarchical partitioning. On an ultrametric tree, use only branch segments within the last
depthunits before the tips. Must not exceed the tree height.NULL(default) uses the current shared mean-depth normaliser. Requireshierarchyand a phylogenetictree.
Value
A long-format data.frame of class hill_dissimilarity (default,
with a plot() method), or a matrix/vector of dissimilarities when
out = "matrix".
Examples
counts <- matrix(c(10, 0, 5, 2, 8, 1), nrow = 3,
                 dimnames = list(c("t1", "t2", "t3"), c("s1", "s2")))
hilldiss(counts)
#> dissimilarity from neutral Hill numbers of "q0", "q1", and "q2".
#> <hilldiv3 result: neutral>
#> 12 rows x 3 cols
#>
#>    q metric     value
#> 1  0      S 0.3333333
#> 2  1      S 0.6081390
#> 3  2      S 0.7308320
#> 4  0      C 0.2000000
#> 5  1      C 0.5229848
#> 6  2      C 0.7308320
#> 7  0      U 0.3333333
#> 8  1      U 0.5229848
#> 9  2      U 0.5758355
#> 10 0      V 0.2000000
#> 11 1      V 0.4369251
#> 12 2      V 0.5758355
plot(hilldiss(counts))
#> dissimilarity from neutral Hill numbers of "q0", "q1", and "q2".